IMDRF issues guidances on cybersecurity, personalized devices, conformity assessment bodies and terminologies for categorized Adverse Event Reporting (AER)
The International Medical Device Regulators Forum (IMDRF) issued on April 20th four technical documents covering cybersecurity, personalized devices, conformity assessment bodies and terminologies for categorized Adverse Event Reporting (AER). Following, a short overview on the content of the guidelines:
Principles and Practices for Medical Device Cybersecurity
“The purpose of this IMDRF guidance document is to provide general principles and best practices to facilitate international regulatory convergence on medical device cybersecurity.”
“This document is designed to provide concrete recommendations to all responsible stakeholders on the general principles and best practices for medical device cybersecurity (including in vitro diagnostic (IVD) medical devices). It outlines recommendations for medical device manufacturers, healthcare providers, regulators, and users to minimize cybersecurity risks that could arise from use of the device for its intended purposes and to ensure maintenance and continuity of device safety and performance.”
Requirements for Regulatory Authority Recognition of Conformity Assessment Bodies (CABs) Conducting Medical Device Regulatory Reviews
“The purpose of this document is to define the requirements for CABs performing regulatory reviews and other related functions for medical devices, including IVD medical devices. Both the regulatory review process and the decisions made by a CAB may be subject to further review by the applicable Regulatory Authority in the countries and regions where the medical device is manufactured and/or placed on the market.”
Personalized Medical Devices – Regulatory Pathways
“The purpose of this IMDRF guidance is to recommend a harmonized approach for the application of existing regulatory pathways to medical devices that are intended for a particular individual, and to identify special considerations for the regulation of each identified category of personalized medical device.”
IMDRF terminologies for categorized Adverse Event Reporting (AER): terms, terminology structure and codes
“This document has been prepared by the IMDRF Adverse Event Working Group, charged with developing a harmonized terminology for reporting adverse events related to medical devices including in-vitro diagnostics (IVDs).”
“This document provides the IMDRF terms, definitions and IMDRF alpha-numerical codes to be used for Adverse Event (AE) reporting concerning medical devices and in vitro diagnostics both pre and post market.”
For more details, please see the original documents on the IMDRF website.
Contact us to learn more
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
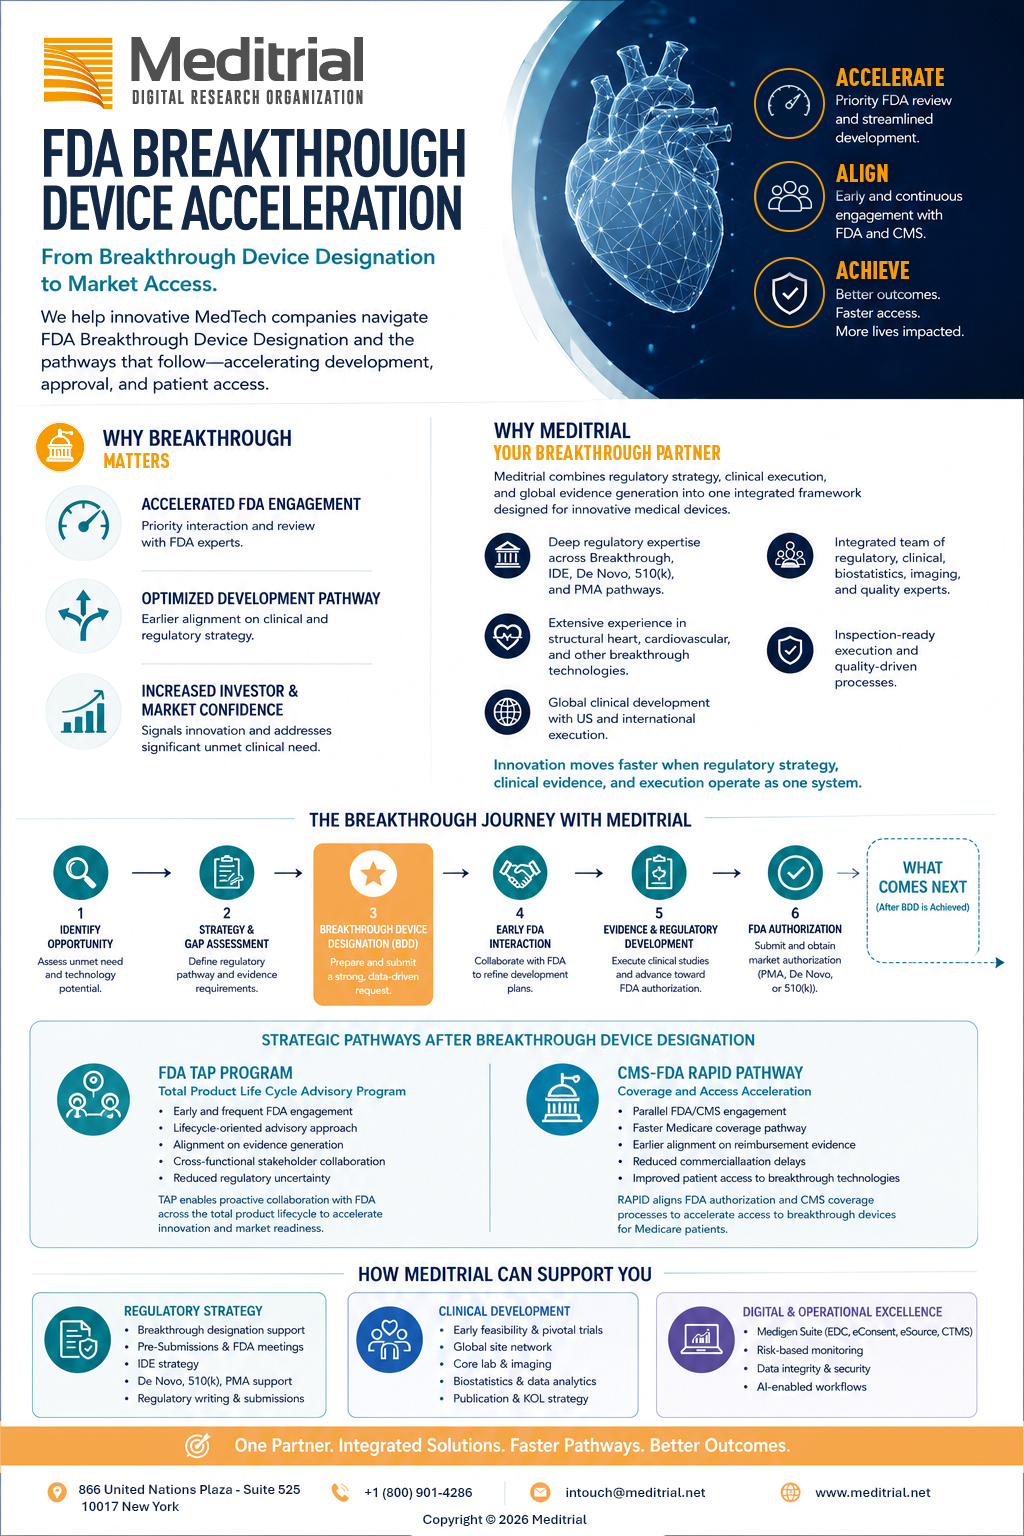
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.